Featured Projects
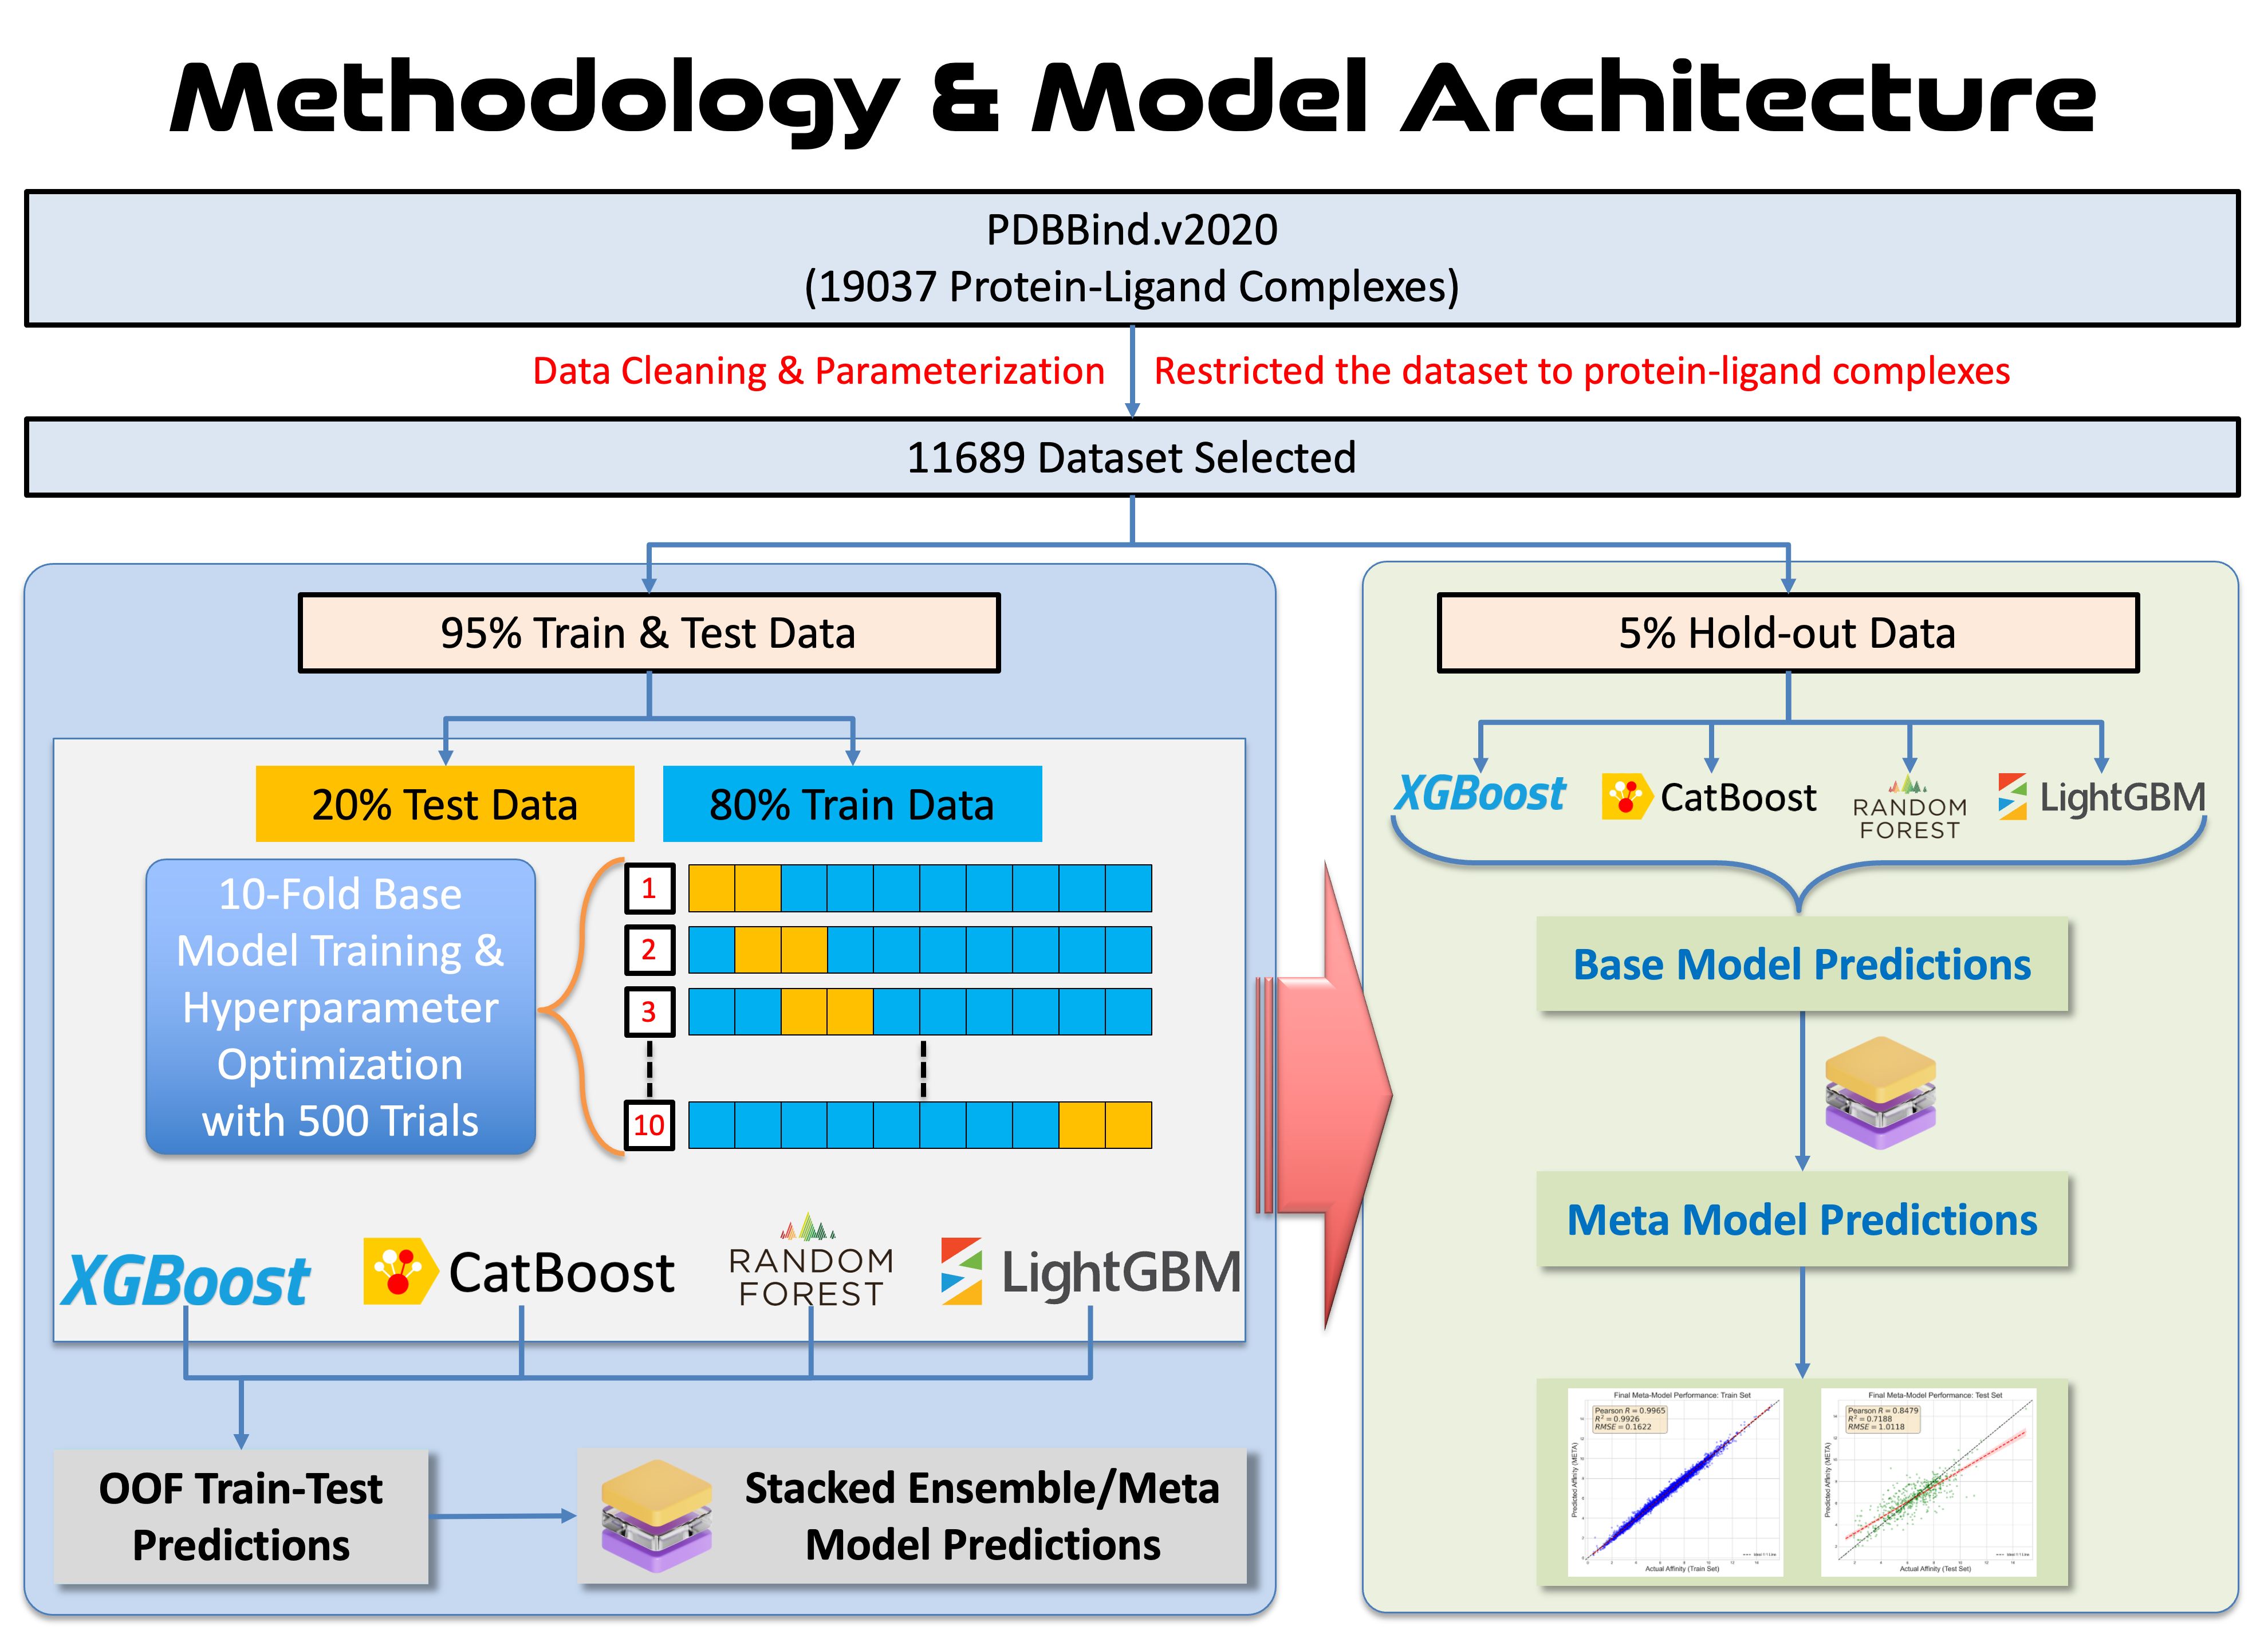
SEARCH-ML Framework
Virtual Screening ToolA comprehensive scoring function developed to improve the accuracy of protein-ligand binding affinity predictions. Trained on a curated dataset of 11,689 complexes from PDBbind, this tool utilizes an ensemble of machine learning algorithms to outperform classical scoring functions.
Python
Scikit-Learn
Bash Scripting
HPC
Read Documentation
ProteoLig-ML
Target IdentificationAn ongoing initiative to map protein-ligand interactions across specific viral targets. This project integrates molecular docking pipelines with machine learning refinement stages to filter false positives in high-throughput screening campaigns.
Bioinformatics
Docking
Web Dev